There are some situations in which even finite elements aren't enough. Finite elements allow us to average and interpolate over very small volumes, approaching "d"x along any chosen coordinate axis. But what if we need more than that? For example, there are many situations in neuroscience that go well beyond the trans-membrane ionic model. One of the best examples is the growth cone of an axon. In an axonal growth cone, filopodia extend out of inner structures called lammelopodia. The lamellopodia are bulbous caps at the end of the growing axons. They are stabilized by a network of actin filaments attached to the cytoskeleton. 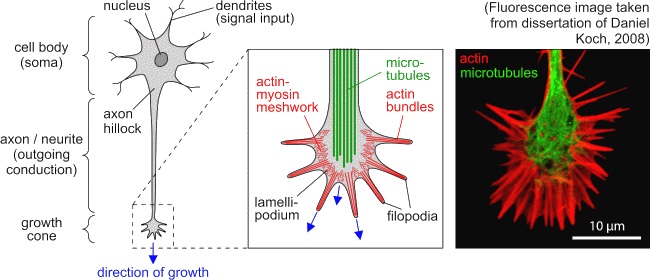 What Is Molecular Dynamics?The prototype use case for molecular dynamics is the diffusion of molecules in the plane of the plasma membrane. This video is a great introduction to molecular dynamics. In addition to the lipid bilayer, biological membranes have some large molecules inserted into them, some are trans-membrane and some are on one side or the other. Phospho-lipids and glyco-lipids play an important role in the membrane, as do molecules like cholesterol. The membrane-bound molecules have a close relationship with the cytoskeleton, which is a network of actin fibers immediately underneath the plasma membrane. Some membrane-bound structures are anchored to the cytoskeleton, while others float around in the fluid mosaic. In a filopodium, the chemical sensors are located at the growing tips, and around them are anchor points for proteins that support actin linkage and growth. But new actin molecules are not synthesized in the tips, they're synthesized in the endoplasmic reticulum and they have to be transported to the tips. Molecular dynamics allows us to visualize these processes. How Molecular Dynamics WorksIn a plasma membrane, the molecules are constrained to move in the plane of the membrane. However in an intracellular volume, this is not the case. If we're interested in ion diffusion from the tip of a growth cone to the base of an actin filament, we have a three dimensional situation where the boundaries are determined by the filopodial membrane. When a calcium ion enters the filopodium through a channel, it can move forward in the direction of the actin fiber, but it can also move sideways and then bind to trans-membrane molecules. The downside of molecular modeling is we don't know 100% of the interactions. There could be molecules there we haven't discovered yet and know nothing about. In such cases, what generally happens is the delta between what we want and what the simulation tells us, is fudged into a lump term without specifying the details. Sometimes this works, and sometimes it doesn't. It worked spectacularly well for Ilya Prigogine, it got him a Nobel prize. But that was back in the 70's, and these days we have higher standards (hopefully). Much of Prigogine's work was done on an IBM-370/158, which was the first IBM model to use semiconductor memory instead of cores. It had 64 Mb maximum memory, based on its 24-bit architecture, and in most cases there was considerably less (the first models offered either half or a full megabyte). How Do We Visualize MD Results?When we visualize molecular dynamics, we want to be able to easily distinguish the molecules. Usually this is done with color. And, we'd like to represent the size and shape of molecules "reasonably accurately", which means our graphics engine can reference reusable primitives as long as they can be rendered reasonably quickly. All we're going to see on an MD screen, is a bunch of colored molecules floating around. If we're interested in something specific like calcium-based actin linkage, we're going to have to highlight that particular activity in a special way to make it visible on the screen. (Otherwise, it'll disappear into a sea of color). We may then be interested in knowing how many molecules are in a particular state at any given time. We have two choices, we can have the simulator output that information up front, or we can count colored molecules from a display. Sometimes counting is easier and faster, it's a "just-in-time" concept that unburdens the simulator from an extra traversal of the mesh.What About The Boundaries?In an MD simulation, molecules react with each other, and whether or not they bounce of walls depends on whether there's any reaction involved. Walls have charge, and so do molecules, and sometimes some stickiness will occur between hydrophilic and hydrophobic domains on that basis. There isn't really a hard-and-fast "boundary" per se in an MD simulation. The properties of the boundary are determined by the properties of the molecules. If you want to add a Debye layer in a finite element simulation, you have to build a special boundary for it, because it responds differently to charge than the rest of the volume. However in an MD simulation the Debye layer forms by itself, you don't have to do anything special, it happens naturally as a result of the physics. This is an important point, because the same thing happens with IAF neurons in a traditional simulator, when you have to specify the threshold instead of letting it determine itself.Is This Really Worth Doing?Even with AI in the loop, an MD simulation is still a lot of work. On the other hand, traditional simulators are toys, they're not going to tell us anything new. Finite elements are in the middle, they're more detailed than cable models and more accessible computationally and visualization-wise, but there's a level of detail they simply won't get to, and sometimes we care about that, and sometimes we don't. Right now at this point in history, we care about things like astrocyte leaflets and ephaptic communication. Tomorrow we may have all that figured out and we'll be trying to manipulate attractors. These days, things are moving so fast that anything you want to research will become a moving target before you're done investigating it. If you decide to roll your own toolset out of need, the chances are good that there will no longer be a need by the time you get done developing it. An MD simulation takes a year, and it's a lot of work. Sometimes the goal is worth it, but you can use your judgement to determine if things will remain stable long enough for you to get the job done. |
Next: Random Numbers